PharmaLectures.com

Courses › GMP, GDP, quality and compliance › URS to PQ: qualification of equipment, utilities and facilities without the paper mountain

URS to PQ: qualification of equipment, utilities and facilities without the paper mountain

Qualification files that pass first review, so your new line starts on schedule.

Equipment qualification is the documented proof that equipment, utilities and facilities used to make medicines are designed, installed and work as intended. In the European Union the rules sit in Annex 15 of the EU GMP guide, revised in 2015, which describes a lifecycle from the user requirements specification through design qualification, installation qualification, operational qualification and performance qualification, often shortened to URS, DQ, IQ, OQ and PQ. Qualified status must then be kept through change control and periodic review. This course takes engineering and QA staff from URS to PQ.

  • 10 audio modulesthree lengths each
  • 1 h 40 min to 8 h 20 minbriefing to expanded
  • Workbook, transcript, certificateand a recorded quiz per module
  • Engineering, QAwho it is written for

Why equipment qualification training matters

The Installation Qualification for a new line misses one utility connection. Quality Assurance rejects the protocol, and later an inspector asks why nobody caught the gap. The line sits idle while engineers rewrite documents instead of running batches.

What changes after this course

Your engineers write User Requirements Specifications that vendors follow, and protocols that Quality Assurance approves on the first read. You hold a certificate per learner, a recorded questionnaire result per module and a training record for each person.

Watch the pitch

3:43 on how an audio course is bought, listened to and recorded. Full screen, one click.

Watch the pitch3:43, full screen, one click

Equipment qualification training: what this course gives your people

A requirement you write today decides the retrofit you avoid next year. Your engineers read EudraLex Volume 4, Chapter 3 and Annex 15 against the actual rooms and utilities, write User Requirements Specifications that vendors cannot reinterpret, and close Design Qualification before the purchase order goes out. The missing parameter gets caught in a meeting, not on the factory floor.

Installation Qualification and Operational Qualification stop coming back with red ink. Your validation specialists record each utility connection, test alarm limits and check safety features while the protocol is still open. Quality Assurance reviews a file with no missing signatures and no half-explained deviations, so handover to manufacturing happens on the planned date.

Calibration intervals get a reason you can say out loud. Your maintenance managers justify each interval and each preventive maintenance task with the equipment's own data, in line with Good Manufacturing Practice. When production asks to modify a machine, your team checks the change control record and answers on the spot whether requalification is needed.

In the audit room, your site leaders open the validation master plan and the qualification reports without searching. A Food and Drug Administration investigator sees the qualified state of each system documented, which lowers the chance of a Form 483 observation on equipment, calibration or maintenance records under 21 Code of Federal Regulations Part 211.

Who needs equipment qualification training

  • Validation Engineer

    Quality Assurance sends your Installation Qualification protocol back for missing test points. You leave knowing which data to capture, so the next protocol is approved on the first read.

  • Quality Assurance Specialist

    You spend days checking a Performance Qualification report against the original requirements. You get a clear order of checks against EudraLex Annex 15 that shortens that review.

  • Facilities Manager

    An auditor asks for your utility calibration schedule and you know the intervals have no written reason behind them. You leave able to build maintenance programs with a justification for each one.

  • Project Engineer

    The vendor delivers a machine that misses your process needs and points to the contract wording. You write the User Requirements Specification so the supplier is held to it before delivery.

Roles: Quality assurance and Qualified Person, Production and engineering

Businesses: every kind of pharma company

Countries: India, China, Türkiye, Korea, Japan, Taiwan, Egypt, Gulf and MENA, Brazil and Latin America, Pakistan, Bangladesh and Vietnam, United States, Europe

Selling into: any jurisdiction

Courses are in English. Other languages on request.

After this course your people can

  1. Write a User Requirements Specification that states exact performance limits and critical process parameters for a vendor to meet.
  2. Score vendor proposals against your requirements and record the result in a Design Qualification review before any purchase order.
  3. Record utility connections and safety features in an Installation Qualification protocol that Quality Assurance can approve without queries.
  4. Set alarm limits and test operating ranges so the Operational Qualification protocol is complete and defensible.
  5. Set calibration intervals from instrument history and write the reason into the maintenance schedule.
  6. Decide from a change control request whether a modification requires requalification, and record the reasoning.
  7. Present a validation master plan and Site Master File to an inspector in the opening meeting.

What the company gets

  • New manufacturing lines stay on schedule because validation protocols stop bouncing between engineering and Quality Assurance.
  • The risk of a Food and Drug Administration Form 483 observation on calibration or missing maintenance records goes down, because every instrument has a documented reason behind its interval.
  • Auditors receive a validation master plan that sets out your qualification approach and scope, plus reports that show the qualified state of each system.
  • You can show who is trained with a per-person training record, a recorded questionnaire per module and a certificate per learner.

The evidence: questionnaire, training record, certificate

EU GMP Chapter 2, section 2.11, and 21 CFR 211.25 both ask for training that is documented and shown to work. This is what each learner leaves behind.

A recorded questionnaire per module

Every one of the 10 modules ends with a multiple-choice questionnaire. The result is recorded per person with a timestamp, whichever of the three lengths the person listened to.

A per-person training record

The export lists each learner, each module, the result and the date, which is the training record EU GMP Chapter 2 and 21 CFR 211.25 ask you to keep and the first thing an inspector asks for.

A certificate per learner

Name, company, course, hours, the guideline version taught and the date. It documents in-company training; it is not a professional qualification or an accreditation.

Your pass mark, your rule

There is no fixed pass mark. Your quality or human-resources manager decides what counts as passed and when a module is retaken, exactly as your training procedure says.

Delivered within 72 hours of payment, with the workbook, the full transcript and a source for every fact in the audio.

How long it takes, by who is listening

Nobody spends a day in a classroom. Every module exists in three lengths; a person takes the length their role needs, and every length ends with the same recorded questionnaire, so the training record is complete either way.

Briefing1 h 40 min10 min a module, for executives, sales and anyone who must know the rules, not run them
Standard4 h 10 min25 min a module, for regulatory, quality and business-development staff
Expanded8 h 20 min50 min a module, for technical staff who run the procedure day to day
Who takes this courseLengthTotal time
Quality assurance and Qualified PersonExpanded8 h 20 min
Production and engineeringExpanded8 h 20 min

Equipment qualification course outline: 10 modules

10 modules. Under each title are the three things the listener can do afterwards; the audio itself runs 10 to 50 minutes depending on the length chosen.

  1. 1Apply Good Manufacturing Practice rules to facility design from day one

    • How to trace a Good Manufacturing Practice rule to the room, pipe or air handler that has to meet it.
    • Why EudraLex Annex 15 treats qualification as a lifecycle, and how that spares you rework after late changes.
    • When a facility upgrade pulls an approved area back into scope, and when it leaves the file alone.
  2. 2Write User Requirements Specifications that vendors cannot misread

    • How to fix critical process parameters in writing before the first vendor quote arrives.
    • The requirement wording that stops a supplier from reading a gap as permission to substitute.
    • What to do when production, quality and engineering each want a different requirement in one document.
  3. 3Score vendors and close Design Qualification before you place the order

    • How to score proposals against your requirements so the cheapest bid cannot win by leaving things out.
    • Why a Design Qualification review before the purchase order costs you a meeting, not a retrofit.
    • When to turn down a supplier whose delivery date is the best you have been offered.
  4. 4Execute Installation Qualification without a single missed connection

    • What to do when the delivered machine does not match the vendor drawings you approved.
    • How to record every utility connection so the Quality Assurance reviewer has nothing to send back.
    • The safety feature check that belongs on your Installation Qualification checklist before anyone signs.
  5. 5Run Operational Qualification that proves the equipment works as specified

    • How to push equipment to its limits in testing without damaging the new components.
    • Why alarm limits set tightly during Operational Qualification spare you deviations in routine production.
    • The test points an inspector looks for when they read your operating ranges.
  6. 6Build Performance Qualification tests that show consistent batch quality

    • How to pick representative materials so the run proves the system handles real production.
    • What to do when a Performance Qualification batch misses a release specification.
    • The test plan that shows consistent quality in the terms EudraLex Annex 15 uses.
  7. 7Set calibration and maintenance schedules that keep your utilities running

    • How to set a calibration interval you can defend with the instrument's own drift history.
    • Why a preventive maintenance schedule built on equipment data keeps utilities off the audit finding list.
    • The records an auditor asks for to see that your instruments read correctly, ready in one folder.
  8. 8Decide when a change triggers requalification, and prove it in writing

    • When a small equipment tweak needs full requalification, and when a documented assessment is enough.
    • How to write a change control request that engineering and quality both sign without a meeting.
    • Why a documented change assessment comes before any decision on requalification.
  9. 9Write validation master plans that Quality Assurance approves fast

    • How to structure a qualification report so an inspector finds the answer in seconds.
    • What to do when you find a deviation halfway through an approved protocol.
    • The signature order that stops anyone changing final data after approval.
  10. 10Answer inspector questions on equipment and facility status on the spot

    • How to explain your validation strategy to an investigator who challenges your method.
    • Why a tidy equipment logbook keeps an auditor from asking to see the next system.
    • The way to present your Site Master File in the opening meeting so the questions stay on topic.
The founders

One of us was inspected. The other one pays for the training.

Every claim below is sourced on the about page: the medicines register, the Lithuanian Pharmaceutical Union board list, the university directory.

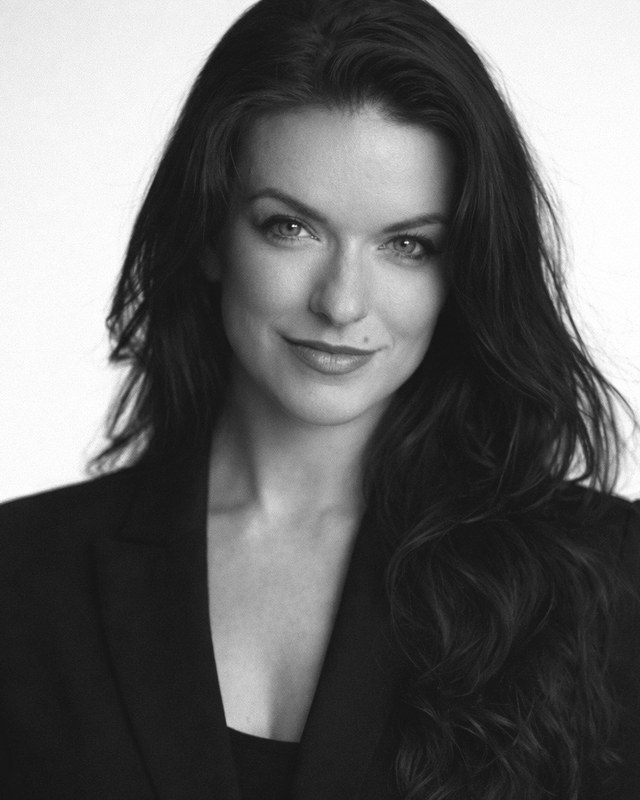
Justina Krukonytė-Racz

Justina Krukonytė-Racz

Co-founder. Licensed pharmacist.

GDPResponsible PersonWarehouse & logistics

Licensed pharmacist. Responsible Person at Tamro (PHOENIX group), where she led the GDP set-up of the distribution centre.

The Responsible Person is the named person the regulator holds accountable for every pallet that leaves the building.

She writes the modules around the procedures a GDP inspection checks. Board member of Lithuanian Pharmaceutical Union.

Listed in the Lithuanian medicines register as Pharmacist.
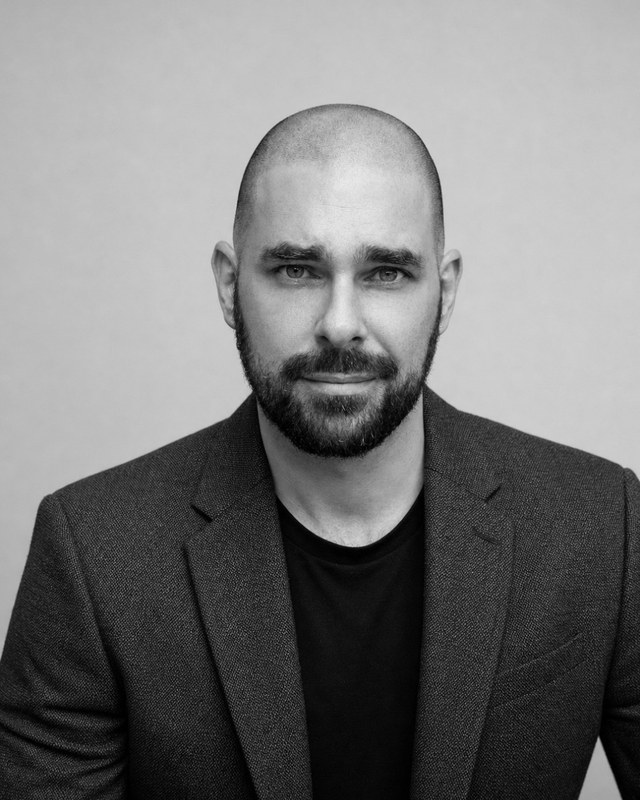
Prof. Ben Racz

Prof. Ben Racz

Co-founder. Builds the platform and is responsible for delivery, records and pricing.

Platform & deliveryQuantitative researchCarnegie Mellon

He was Professor of the Practice at the College of Engineering at Carnegie Mellon University, where he taught Advanced Quantitative Finance and Algorithmic Trading to MSc students.

He is the founder and Managing Director of Algorithmic Sciences, a scientific research and consulting firm. Earlier he co-founded a data-driven growth consultancy, a data-science based wholesale trading company and an information security firm.

More about Ben at benracz.com

Questions about equipment qualification training

What is the difference between IQ, OQ and PQ?

Installation qualification checks that equipment is delivered and installed as specified, with the right parts, connections, drawings and manuals. Operational qualification checks that it operates as intended across its working ranges, including alarms and limits. Performance qualification checks that it consistently performs as required with real or representative materials under normal conditions. EU GMP Annex 15 describes all three as stages of one lifecycle.

When is requalification required?

EU GMP Annex 15 says equipment, facilities, utilities and systems should be evaluated at an appropriate frequency to confirm they remain in a state of control. A change that could affect product quality, assessed through change control, can also trigger requalification of the affected parts. Teams that document the rationale for each decision avoid both needless rework and inspection findings.

Is there a certificate?

Yes. Every learner who finishes the course receives a certificate in their own name, and every module ends with a recorded multiple-choice questionnaire, so the certificate sits on top of a per-person training record. The certificate documents in-company training; it is not a professional qualification or an accreditation, and your quality or human-resources manager sets the pass mark and decides when a module is retaken.

How long does the course take?

10 modules, each in three lengths: a 10-minute briefing, a 25-minute standard run and a 50-minute expanded walk-through. The whole course takes 1 h 40 min in the briefing length and 8 h 20 min in the expanded length. People take the length their role needs, and every length ends with the same recorded questionnaire, so the training record is complete either way.

Who should take this course?

It is written for Engineering, QA. Owners, managers and sales staff hear the briefing length, regulatory, quality and business-development staff take the standard run, and the people who run the procedure every day get the expanded version. One licence covers one named person on this course for a year, so the same title can be rolled out across a whole team at once.

Is this training accepted as a GMP training record by an auditor?

EU GMP Chapter 2 and 21 CFR 211.25 require documented, effective training, and that is what the course produces: a recorded multiple-choice result per module and per person, a certificate per learner and a transcript of what was taught. No provider can promise acceptance by a specific inspector; the record is built to answer what inspectors ask for, and your quality manager decides the pass mark.

How is it delivered and how fast?

The course arrives as audio files with a companion workbook, a full transcript and the questionnaire per module, within 72 hours of your payment arriving. You host the files on your own system or share them as you like; there is no platform to log into and no learner management system to set up. The free sample module arrives on request, before any purchase.

What does it cost per learner?

The list price is 9 euros per module, per learner, per year, so this course is 90 euros for one person for twelve months. The price per learner falls as more people take the same course: 10 learners pay 10 percent less and the discount keeps growing to 50 percent at 1000 learners. Type a number on the price panel and the yearly total appears.

Get this course for your team

One licence is one person on this course for a year. All three lengths, the workbook, the certificate and the training record are included in every purchase. Pick how many people, add it to the cart, and you have an invoice in your inbox.

Free sample module

Hear one full module before you spend a cent. Send your company and the role you want trained, and a complete sample module comes back with its workbook and transcript.

Connect on WhatsApp

or email [email protected], or ask for a written quote for your team